Medicines
SmPC of the 08/06/2026
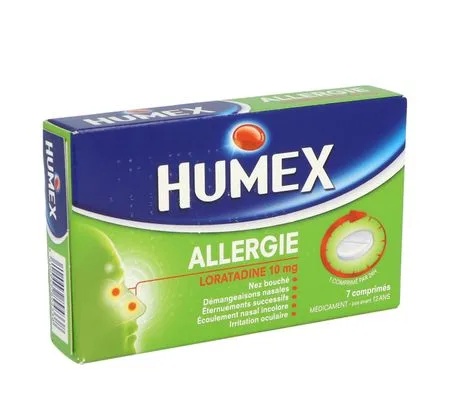
HUMEX ALLERGY LORATADINE 10 mg, tablet
Forms and presentations
Tablet.
White, round tablet with a breaking bar.
The break bar is only there to facilitate taking the tablet; it does not divide it into two equal half-doses.
Composition
Loratadine...................................................................................................................................................... 10 mg
For one tablet.
Excipient with known effect: Each tablet contains 75 mg of lactose monohydrate.
Lactose monohydrate, microcrystalline cellulose, maize starch, magnesium stearate.
Indications
HUMEX ALLERGY LORATADINE 10 mg, tablet is indicated for the symptomatic treatment of allergic rhinitis.
Dosage and administration
Dosage
Adults and children over 12 years : 10 mg once a day, i.e. one tablet once a day.
The tablet may be taken regardless of meals.
Children with a body weight of more than 30 kg :
10 mg once a day, i.e. one tablet once a day.
The 10 mg tablet form is not suitable for children weighing less than 30 kg.
The efficacy and safety of HUMEX ALLERGY LORATADINE have not been established in children under 2 years of age.
Severe hepatic impairment :
In patients with severe hepatic impairment, the initial dose should be reduced due to a risk of reduced clearance of loratadine. An initial dose of 10 mg every two days is recommended for adults and children weighing more than 30 kg.
Elderly subjects or renal impairment :
No dose adjustment is necessary in elderly patients or patients with renal impairment.
Duration of treatment
The duration of treatment should not exceed 3 days without medical advice.
Contraindications
HUMEX ALLERGY LORATADINE is contraindicated in cases of hypersensitivity to loratadine or to any of the excipients of the medicinal product.
Warnings and precautions
HUMEX ALLERGY LORATADINE should be used with caution in patients with severe hepatic impairment (see section Dosage and administration.).
This medicine contains lactose. Patients with galactose intolerance, total lactase deficiency or glucose-galactose malabsorption syndrome (rare hereditary diseases) must not take this medicine.
Administration of HUMEX ALLERGY LORATADINE must be discontinued at least 48 hours before skin tests are performed for the diagnosis of allergy, as antihistamines may inhibit or reduce the skin response.
Interactions
Psychomotor performance studies have not shown potentiation of the effects of HUMEX ALLERGY LORATADINE when administered simultaneously with alcohol.
The risk of interactions with inhibitors of the cytochromes CYP3A4 or CYP2D6 resulting in increased plasma concentrations of loratadine may increase the risk of occurrence of adverse effects (see section Pharmacokinetics).
Fertility, pregnancy and breastfeeding
Pregnancy
Animal studies have not revealed a teratogenic effect of loratadine. The safety of loratadine during pregnancy has not been established. Consequently, the use of HUMEX ALLERGIE LORATADINE during pregnancy is not recommended.
Breast-feeding
Loratadine is excreted in breast milk. Consequently, administration of loratadine during breastfeeding is not recommended.
Driving and using machines
In clinical studies evaluating the ability to drive vehicles, no deleterious effect was observed in patients receiving loratadine. However, patients must be informed that very rarely in certain people somnolence has been described which could affect their ability to drive vehicles or use machines.
Adverse effects
In clinical studies conducted in the paediatric population, in children aged 2 to 12 years, the frequent adverse effects reported with a higher frequency than under placebo were: headache (2,7 %), nervousness (2,3 %) and fatigue (1 %).
In clinical studies conducted in adults and adolescents in the indications allergic rhinitis and chronic idiopathic urticaria, at the recommended dose of 10 mg, adverse effects with loratadine were reported in 2 % more patients than those treated with placebo. The adverse effects most frequently reported with a higher frequency than under placebo were: somnolence (1,2 %), headache (0,6 %), increased appetite (0,5 %) and insomnia (0,1 %).
Other adverse effects very rarely reported since marketing are listed in the following table:
|
Immune system disorders Nervous system disorders Cardiac disorders Gastrointestinal disorders Hepatobiliary disorders Skin and subcutaneous tissue disorders General disorders and administration site conditions Investigations |
Anaphylaxis Vertigo Tachycardia, palpitations Nausea, dry mouth, gastritis Liver function disorders Rash, alopecia Fatigue
Weight gain (frequency not known) |
Reporting of suspected adverse reactions
Reporting suspected adverse reactions after authorisation of the medicinal product is important. It allows continued monitoring of the benefit/risk balance of the medicinal product. Healthcare professionals are asked to report any suspected adverse reaction via the national reporting system: National Agency for the Safety of Medicines and Health Products (ANSM) and the network of Regional Centres of Pharmacovigilance - Website: https://signalement.social-sante.gouv.fr/.
Overdose
Overdose of loratadine increases the occurrence of anticholinergic symptoms. Somnolence, tachycardia and headache have been reported in cases of overdose.
In case of overdose, symptomatic treatment and maintenance of vital functions are recommended. Activated charcoal in suspension in water may possibly be administered. Gastric lavage may be considered.
Loratadine is not eliminated by haemodialysis and it is not known whether peritoneal dialysis eliminates it.
The patient must remain under medical supervision after emergency treatment.
Pharmacodynamics
Pharmacotherapeutic class: Antihistamines for systemic use, ATC code: R06AX13.
Loratadine, the active ingredient of HUMEX ALLERGIE LORATADINE, is a tricyclic antihistamine acting selectively on peripheral H1 receptors.
Loratadine does not exert a significant sedative or anticholinergic effect in the majority of the population when used at the recommended dose.
During long-term treatment, no clinically significant changes in vital functions, biological parameters, clinical examination or electrocardiographic tracings have been observed.
Loratadine has no significant action at H2 receptors. It does not inhibit noradrenaline reuptake and has practically no influence on cardiovascular functions or on intrinsic pacemaker activity.
Pharmacokinetics
After oral administration, loratadine is rapidly and well absorbed and undergoes a significant first-pass hepatic effect, being essentially metabolised by CYP3A4 and CYP2D6. The main metabolite - desloratadine - is pharmacologically active and largely responsible for the clinical effect. Maximum plasma concentrations of loratadine and desloratadine are reached (Tmax) between 1-1,5 hours and 1,5-3,7 hours after administration, respectively.
In controlled clinical studies, an increase in plasma concentrations of loratadine has been reported when administered concomitantly with ketoconazole, erythromycin or cimetidine, but without significant clinical consequence (no change in ECG tracings).
The binding of loratadine to circulating proteins is high (97% to 99%), whereas that of the metabolite is more moderate (73% to 76%).
In healthy volunteers, the distribution half-lives of loratadine and its active metabolite are approximately 1 and 2 hours respectively. The main elimination half-life in healthy volunteers was 8,4 hours (range 3 to 20 hours) for loratadine and 28 hours (range 8,8 to 92 hours) for the main active metabolite.
Approximately 40% of the dose is excreted in urine and 42% in faeces over a period of 10 days, mainly as conjugated metabolites. Approximately 27% of the dose is eliminated in urine during the first 24 hours. Less than 1% of the active substance is excreted in the unchanged active form, loratadine or desloratadine.
The bioavailability of loratadine and its active metabolite is dose-dependent.
The pharmacokinetic profile of loratadine and its metabolites is comparable in healthy adult and elderly volunteers.
Concomitant food intake may cause a slight delay in the absorption of loratadine, without consequence for the clinical effect.
In patients with chronic renal failure, the AUC and peak plasma concentrations (Cmax) of loratadine and its metabolite were higher than the AUC and plasma peaks (Cmax) observed in patients with normal renal function. The mean elimination half-lives of loratadine and its metabolite were not significantly different from those observed in normal subjects. Haemodialysis has no effect on the pharmacokinetics of loratadine and its active metabolite in patients with chronic renal failure.
In patients with chronic hepatic impairment of alcoholic origin, the AUC and peak plasma concentrations (Cmax) of loratadine observed were doubled, whereas the pharmacokinetic profile of the active metabolite was not significantly altered compared with that of patients with normal hepatic function. The elimination half-lives of loratadine and its metabolite were 24 hours and 37 hours respectively, and they increased in parallel with the severity of the hepatic impairment.
Loratadine and its active metabolite are excreted in breast milk in breastfeeding women.
Preclinical safety
Preclinical data reveal no specific risk for humans in the light of conventional safety, pharmacology, repeated-dose toxicity, genotoxicity or carcinogenicity studies.
Studies of reproductive function revealed no teratogenic effect in animals. However, prolonged parturition and reduced viability of offspring were observed in rats exposed to plasma levels (AUC) 10 times higher than those achieved with the doses used clinically.
Prescription, supply and reimbursement
| Marketing authorisation (AMM) |
|
12 RUE DAUPHINE
21000 DIJON
Tel: 03 80 54 50 00 https://urgo.fr/
Source: VIDAL · Updated on 10/11/2026. This information does not replace advice from a healthcare professional.